- Solutions
- Product
- Orion® Molecular Design Platform
- Free Energy
Binding Free Energy Predictions - Accurate, Fast, and Scalable
Need to screen hundreds to thousands of ligands for your lead optimization projects in hours, not weeks? OpenEye provides efficient and effective solutions for estimating the binding free energies of protein–ligand complexes.
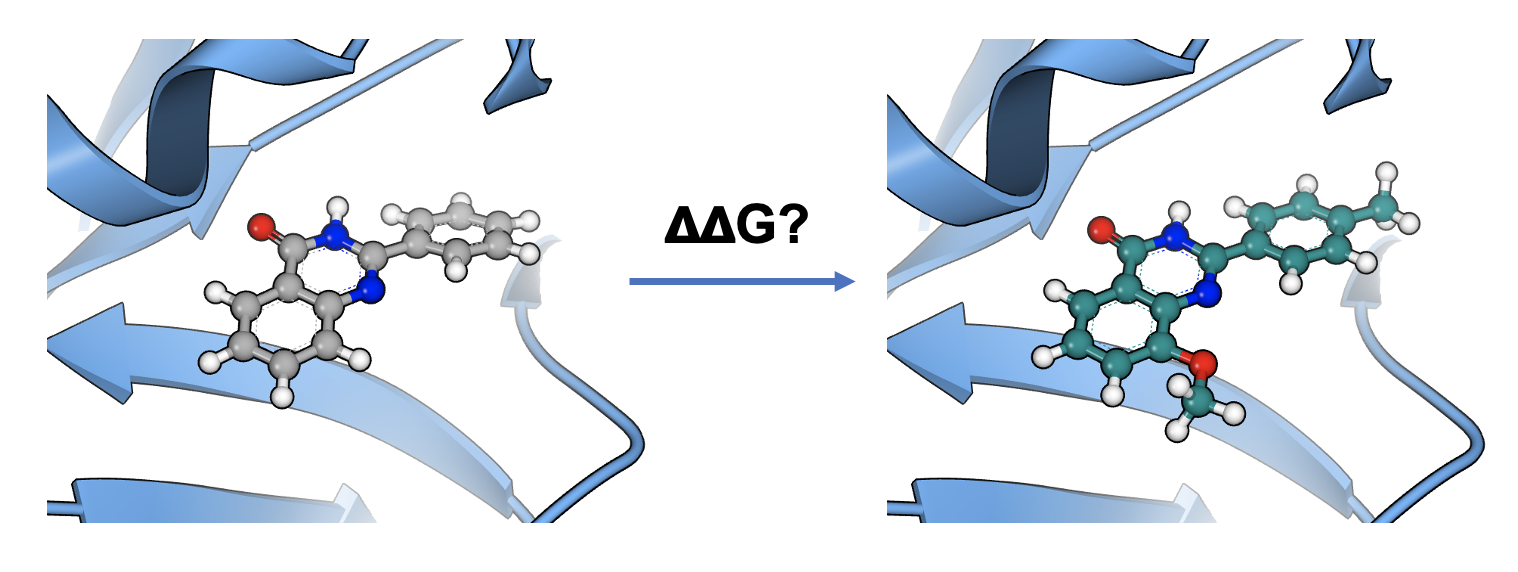
Free Energy Nonequilibrium Switching (FE-NES) calculates ligand–protein binding free energies using the nonequilibrium switching technique pioneered by Gapsys and De Groot (2020) at the Max Planck Institute. By eliminating the extensive equilibrium sampling required by traditional methods like free energy perturbation (FEP), OpenEye’s scientific and technical enhancements make FE-NES significantly faster and with higher throughput. Implemented on the Orion® molecular design platform as an automated, single-step workflow, FE-NES can evaluate hundreds to thousands of ligands in just hours, making it ideal for iterative, real-time lead optimization.
OpenEye’s automated FE-NES solution delivers best-in-class scalability with accuracy comparable to traditional FEP methods, allowing project teams to make confident design decisions without the typical computational bottleneck.
"... 40 ligands could take 24-36 hours with FEP+ but hundreds of ligands with FE-NES only took hours on Orion" - Scientist, CADD, Large Pharma.
Features
- Accurate. Validated on internal and external datasets.
- Fast and Scalable. Highly parallelizable architecture runs hundreds to thousands of ligands in just a few hours.
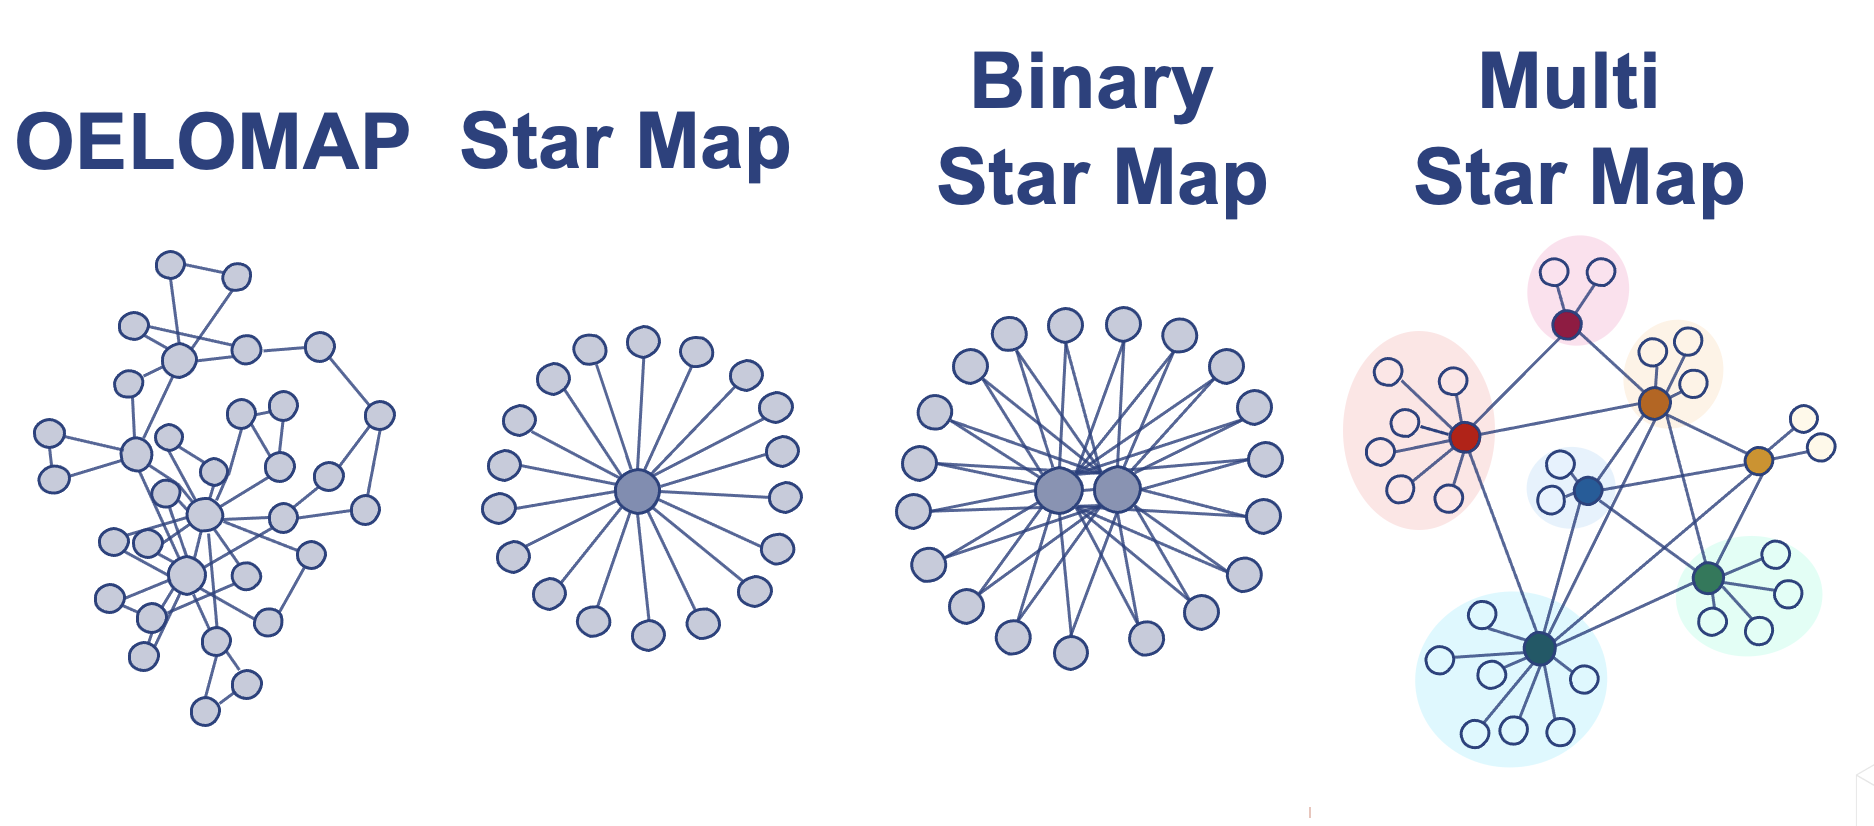
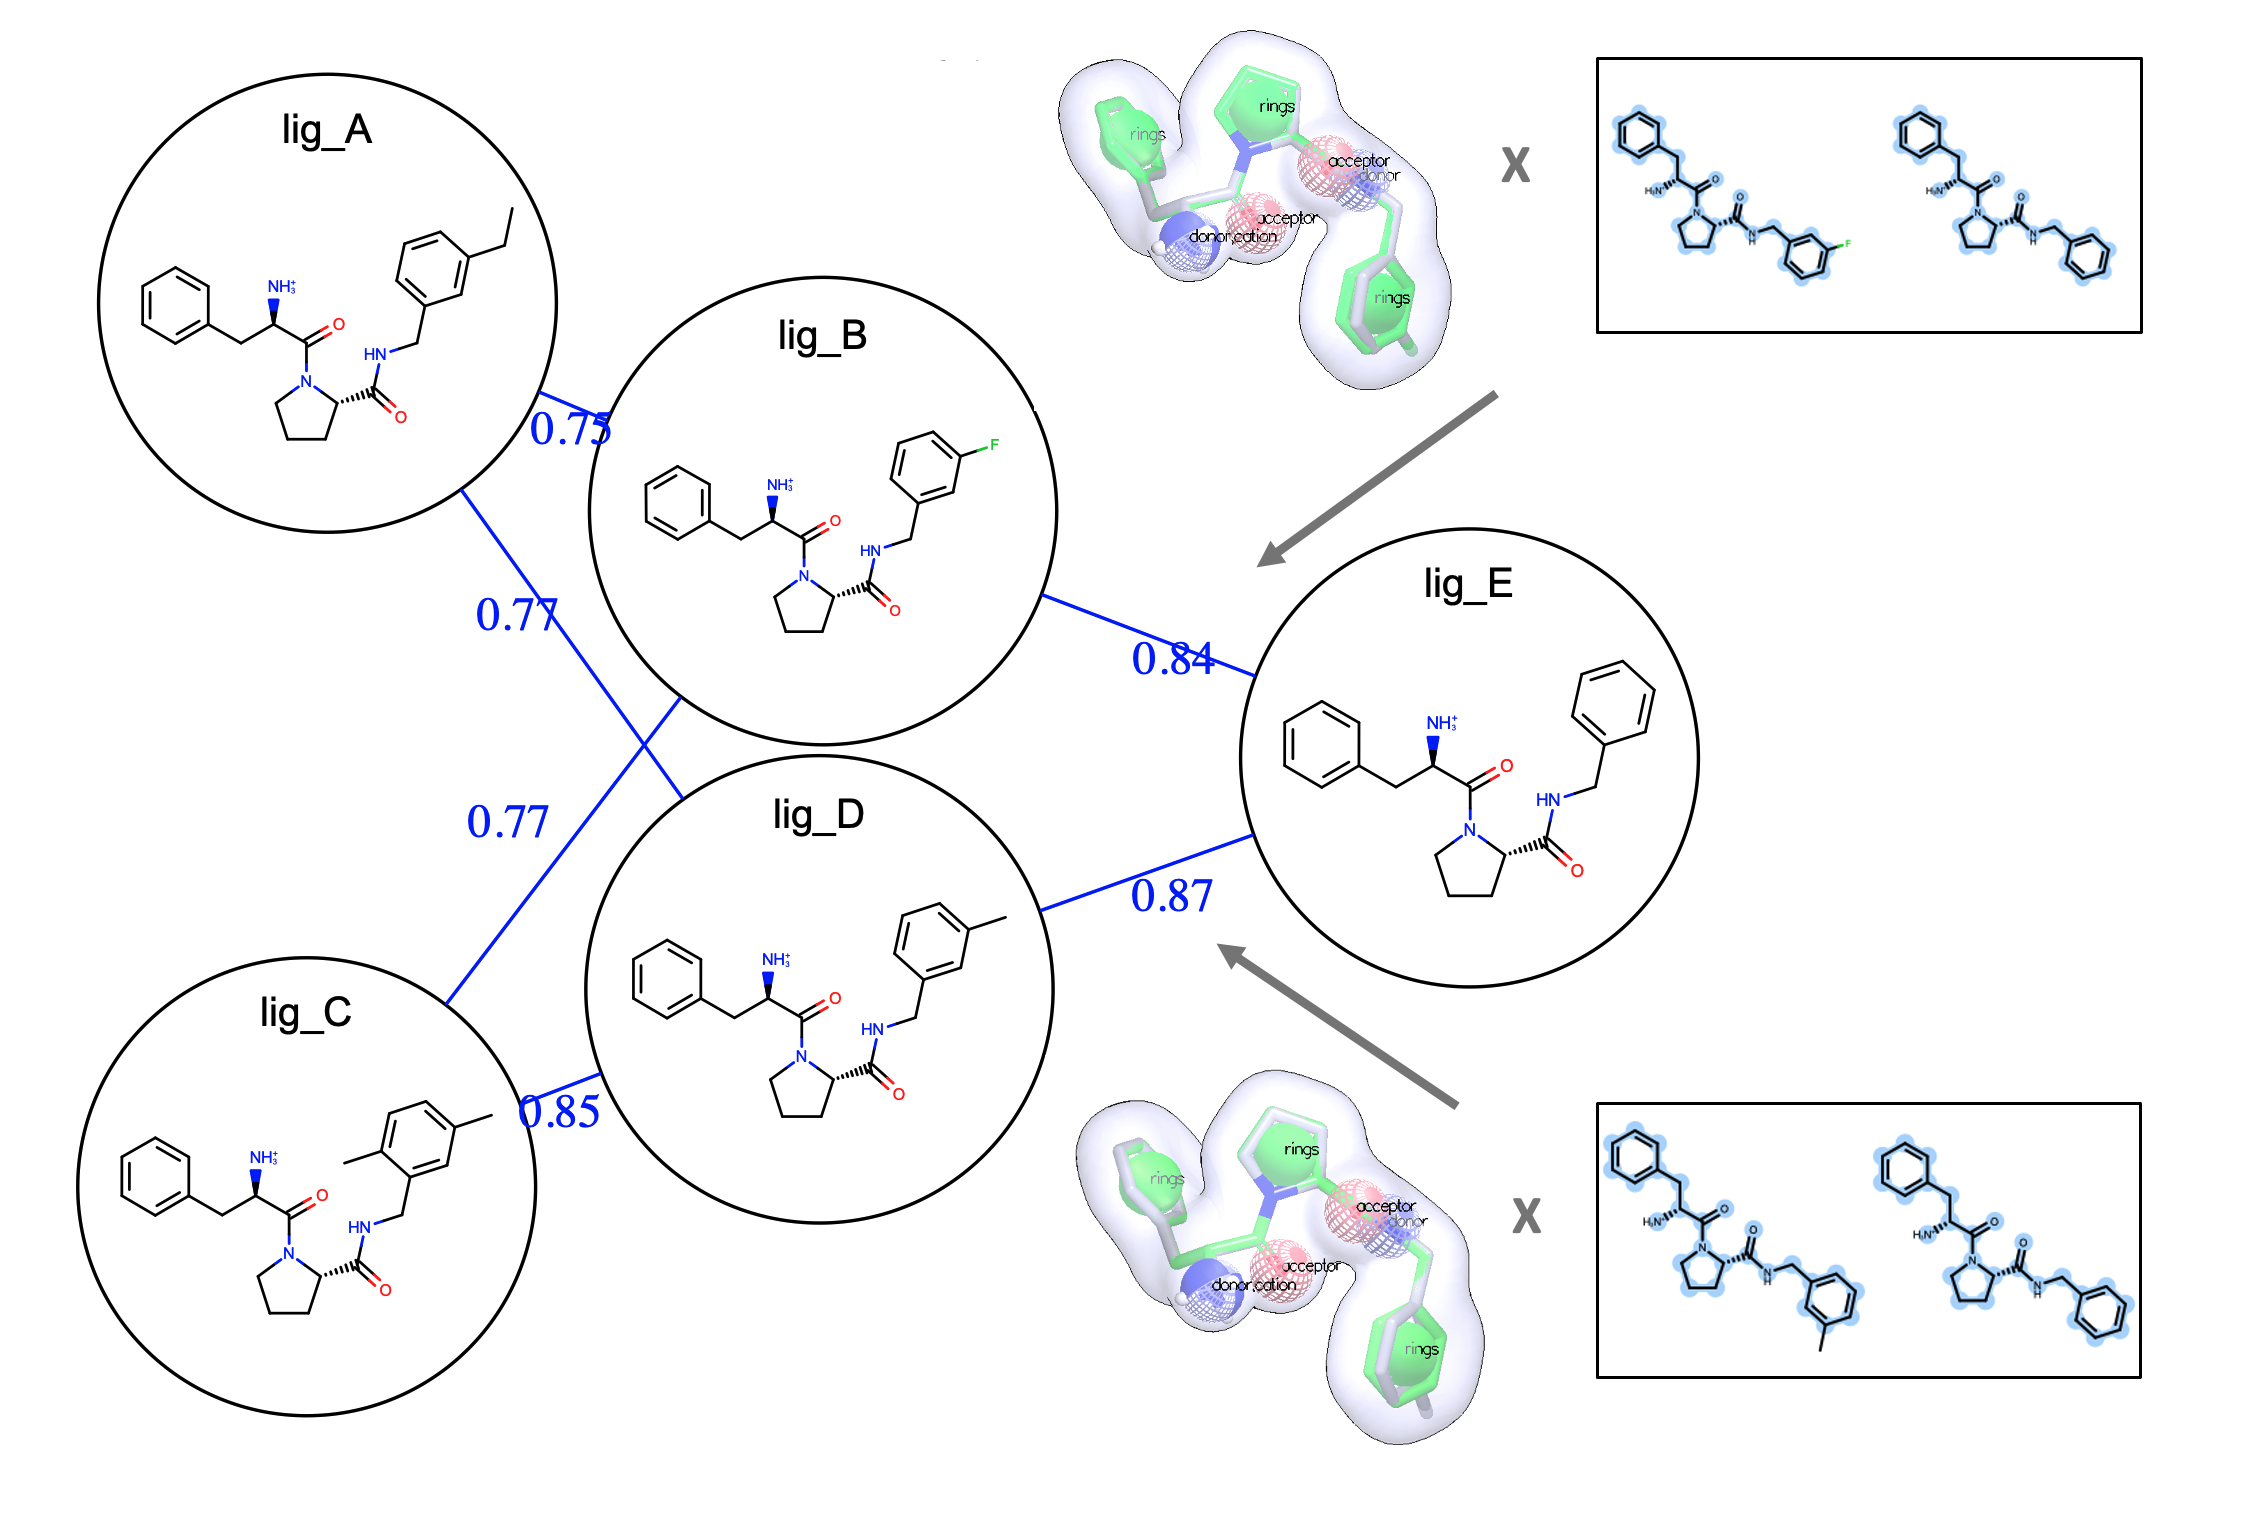
- Flexible. Edge mapper options for connecting ligands include star, binary star, multi star, and OELOMAP with cycle closure.
- Easy-to-Use. Quickly get up and running in a familiar web-based environment.
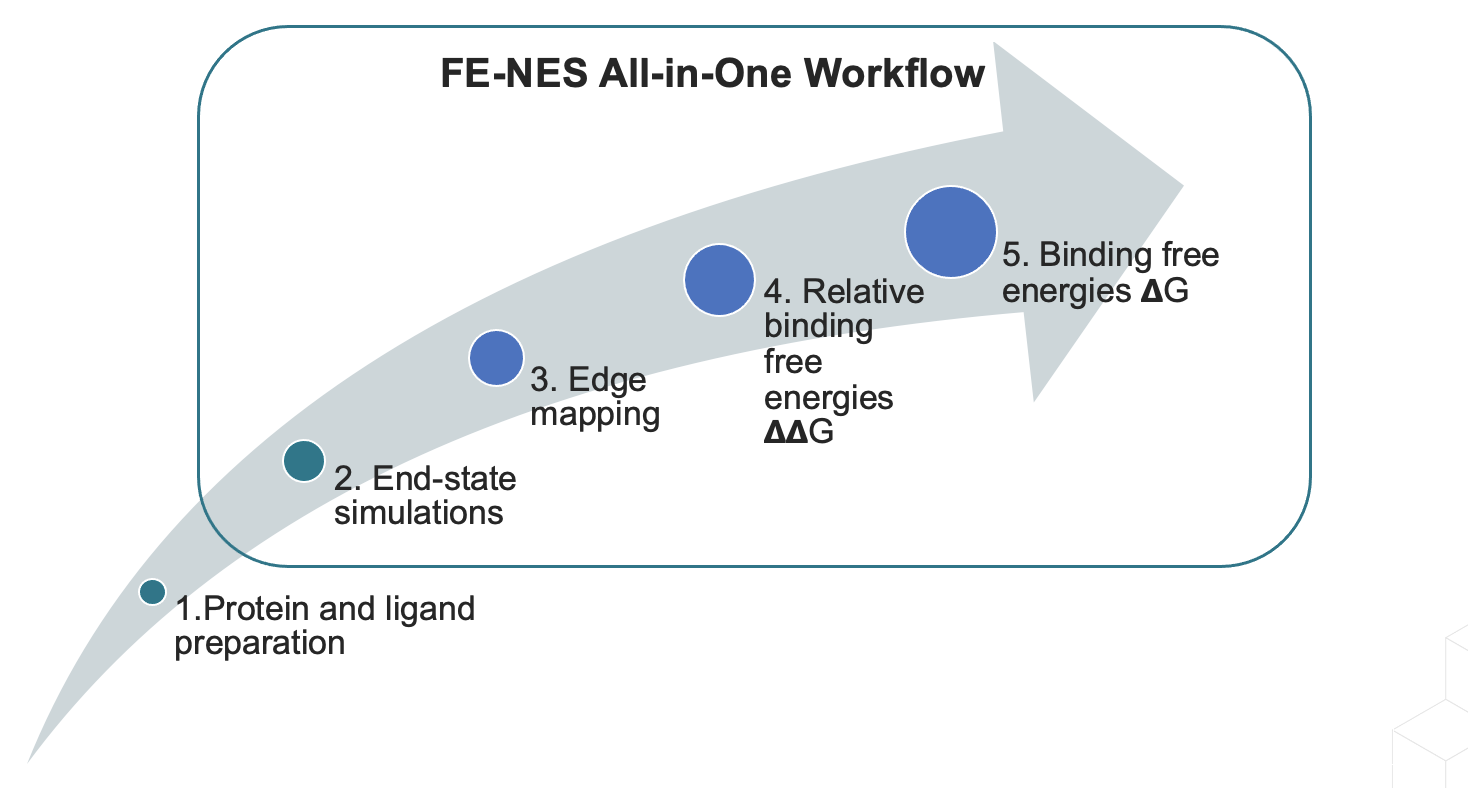
- Automated. Simple all-in-one step process from system set-up, mapping, end-point equilibration, and free energy calculation.
- Control. Highly customizable for novice use and expert control.
- Integrated. Easily combine with other AI and ligand- and structure-based methods.
Rooted in Science. Optimized for Performance.
Evaluate hundreds to thousands of ligands in just hours. FE-NES streamlines drug discovery by bringing powerful, high-throughput free energy calculations straight to your browser.
-
Zero Installation: No complex local setup, access everything securely through your web browser.
-
Maximum Efficiency: Delivering speed, scale, and accuracy to help you optimize leads in real time to meet tight drug discovery project timelines.
Accurately Ranks Ligands by Relative Binding Free Energy
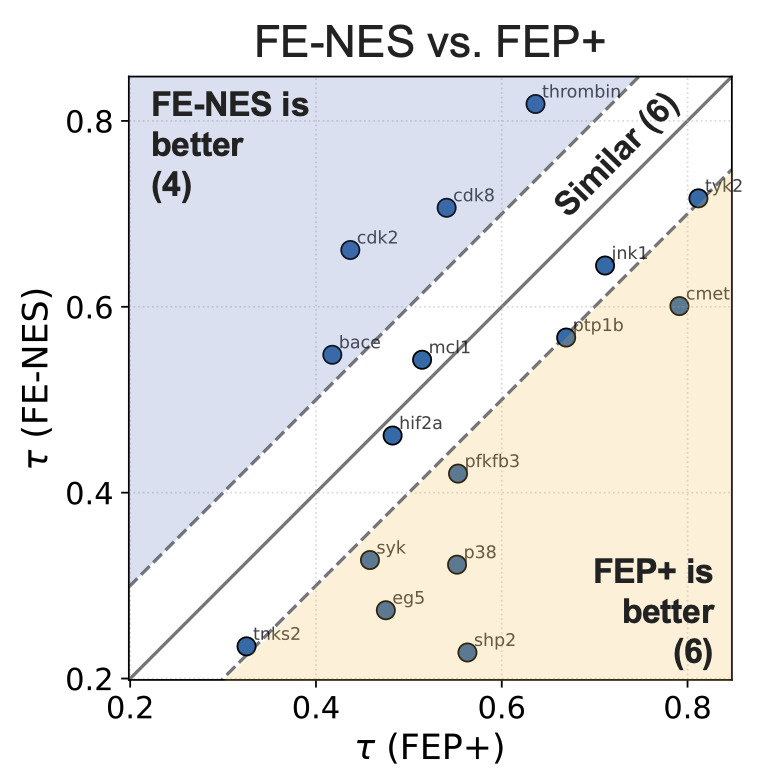
FE-NES accurately ranks ligands by their relative binding free energies to accelerate lead optimization. Kendall’s tau associations show that OpenEye's FE-NES method delivers accuracy comparable to other commercial and open-source approaches. Using the Schindler (2020) and Wang (2015) benchmark datasets from the literature, results demonstrate that FE-NES shows no significant differences in aggregate performance, making it a reliable solution for free energy calculations, with the added benefits of greater speed and scalability.
Datasets: 1) Wang et al., 2015, J. Am. Chem. Soc., 137: 2695. and 2) Schindler et al., 2020, J. Chem. Inf. Model., 60: 5457
Easy, Efficient, and Automated Mapping
With FE-NES, edge mapping now runs automatically as part of the Floe while giving users full flexibility to run it separately if preferred. Users can easily customize their maps using a variety of mapper options, including OELOMAP, Star Map, Binary Star Map, and Multi-Star Map, to connect ligands seamlessly for binding free energy calculations.
Offering Flexibility with the Interactive Edge Mapper
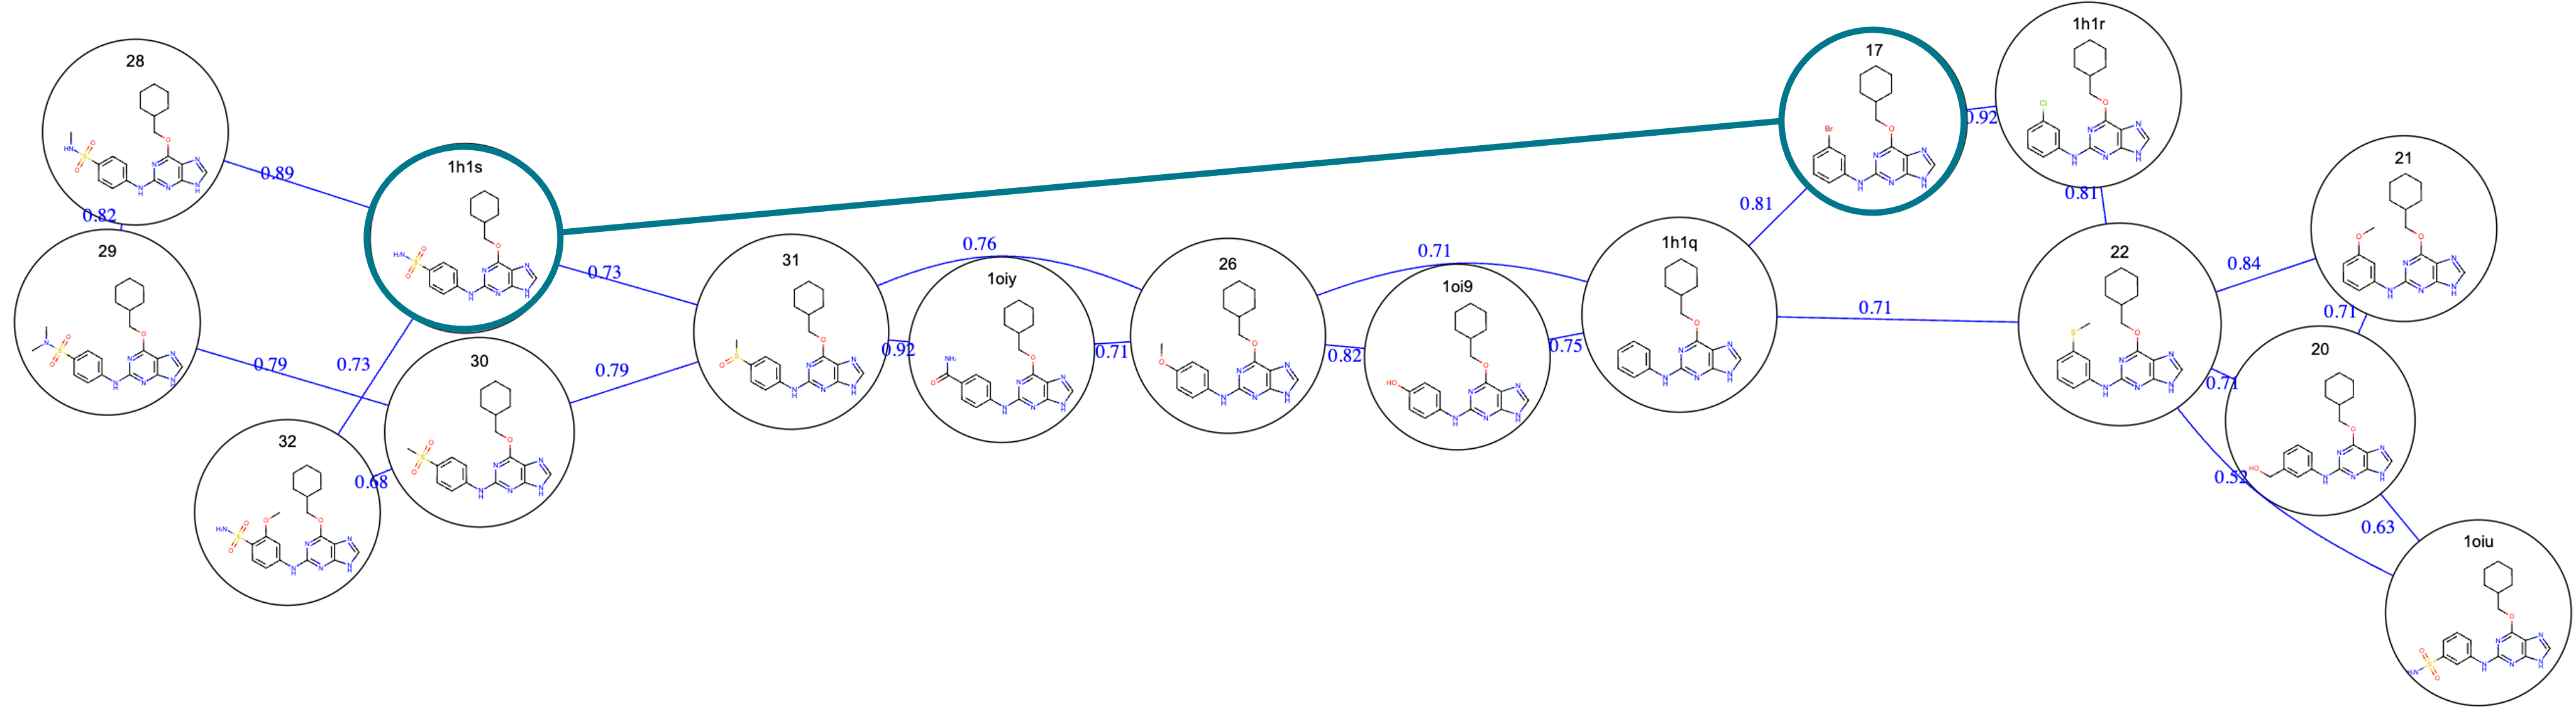
Designed to give scientists fine-grained control over smaller, focused ligand sets, the FE-NES interactive edge mapper allows users to effortlessly add or remove individual edges and re-map selected compounds. The resulting tailored network can be saved directly for downstream calculations, blending algorithmic efficiency with expert structural insight.
Improving Accuracy for Proteins with Flexible Binding Sites
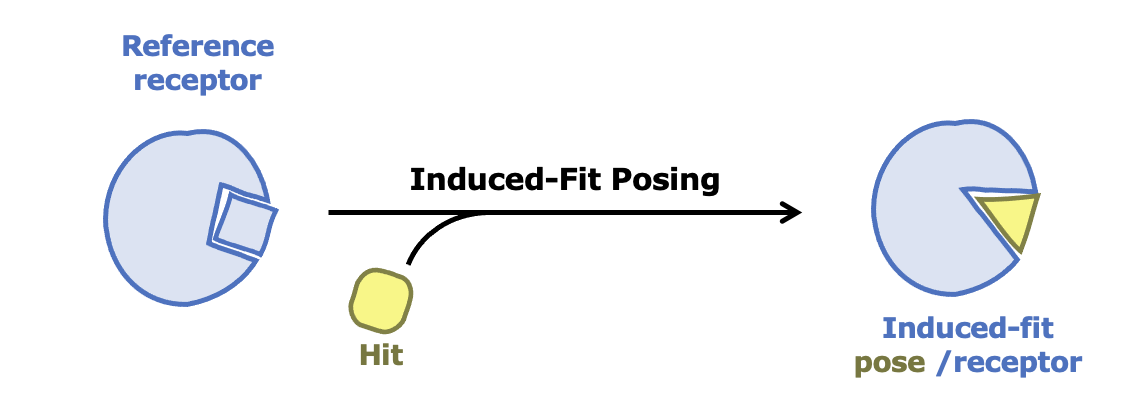
Drug targets sometimes involve proteins with flexible binding sites, which can pose a challenge for accurate modeling. To improve predictions of ligand binding affinities in such cases, combine Induced-Fit Posing (IFP) with Free Energy Nonequilibrium Switching (FE-NES). Together, these tools deliver more accurate and reliable models of ligand–protein interactions.
Learn more about Induced-Fit Posing
FAQs
-
Yes, FE-NES supports alchemical changes between ligands of different formal charges.
-
FE-NES offers versatile workflows for a personalized experience, with options for both novice and expert users. Choose from pre-configured automated workflows or individual workflows for customized tasks with expert control.
-
Users get a comprehensive and interactive report of binding energy calculations. In addition, users can also seamlessly compare experimental affinity with the predicted relative binding free energies.
-
FE-NES offers fast, affordable free energy calculations using a non-equilibrium switching approach, evaluating hundreds to thousands of ligands in just hours.
"... 40 ligands could take 24-36 hours with FEP+ but hundreds of ligands with FE-NES only took hours on Orion" - Scientist, CADD, Large Pharma.
-
FE-NES, a highly efficient and cost-effective free energy calculation method, is adapted from the NES approach advanced by Vytautas Gapsys in the Bert L. DeGroot Lab at the Max Planck Institute for Biophysical Chemistry. (Gapsys et al., Chem. Sci. 2020, 11, 1140-1152). The method is highly parallelizable making it suitable for high throughput calculations.
With the integration of proprietary OpenEye methods for system pre-equilibration and chimeric-molecule calculation, it delivers quick transitions through intermediate states and parallel sampling of short trajectories. This results in very fast, yet equally accurate calculations compared to conventional equilibrium-based methods.
-
FE-NES implements a Maximum Likelihood Estimator to generate per-ligand binding affinities based on predicted relative binding affinities. These per-ligand values will be compared to reference experimental binding free energies if provided.
Learn More
FE-NES helps you rapidly and accurately estimate relative binding free energies, accelerating your drug design process by accurately ranking ligands to guide lead optimization.
For science details, access recording from OpenEye’s miniWebinar "Statistical Considerations in Lead Optimization" with Chris Neale, Ph.D., and "From Prediction to Decision: Streamlining Binding Affinity Workflows" with Dr. Hyesu Jang, Ph.D.
You may also be interested in Science Brief on "Scaling Lead Optimization to Thousands of Compounds. Active Learning with FE-NES and 3D-QSAR" and on "Binding Free Energy Redefined: Accurate, Fast, and Cost Effective".
References
-
Large scale relative protein ligand binding affinities using non-equilibrium alchemy, Gapsys et al., Chemical Science, 2020, 11, 1140-1152
-
Calculation of binding free energies, Gapsys et al., Methods Mol Biol., 2015, 1215:173-209
-
Accurate and reliable prediction of relative ligand binding potency in prospective drug discovery by way of a modern free-energy calculation protocol and force field. Wang et al., J Am Chem Soc. 2015 Feb 25;137(7):2695-703
-
Large-Scale Assessment of Binding Free Energy Calculations in Active Drug Discovery Projects. Schindler et al., J Chem Inf Model. 2020 Nov 23;60(11):5457-5474.
Webinar: Getting More From Your Cryo-EM Data
Webinar: Is the era of "I can't code" over? A chemists' guide to coding with agents.
Webinar: Target X: An Unobstructed View of Pockets
Webinar: Improving the Core: Not Resting on Our Laurels
Webinar: Too Hot, Too Cold, or Past Midnight? Statistical Considerations in Lead Optimization from Goldilocks & Cinderella
Webinar: AI For Drug Discovery
Resources
View Our Recent Webinars
On Demand Webinars
Webinar: From Prediction to Decision: Streamlining Binding Affinity Workflows
Upcoming Webinar
Webinar: Getting More From Your Cryo-EM Data